A mouse study links mitochondrial citrate metabolism to senescent-cell inflammation. Read how the pathway works and what evidence is still needed.


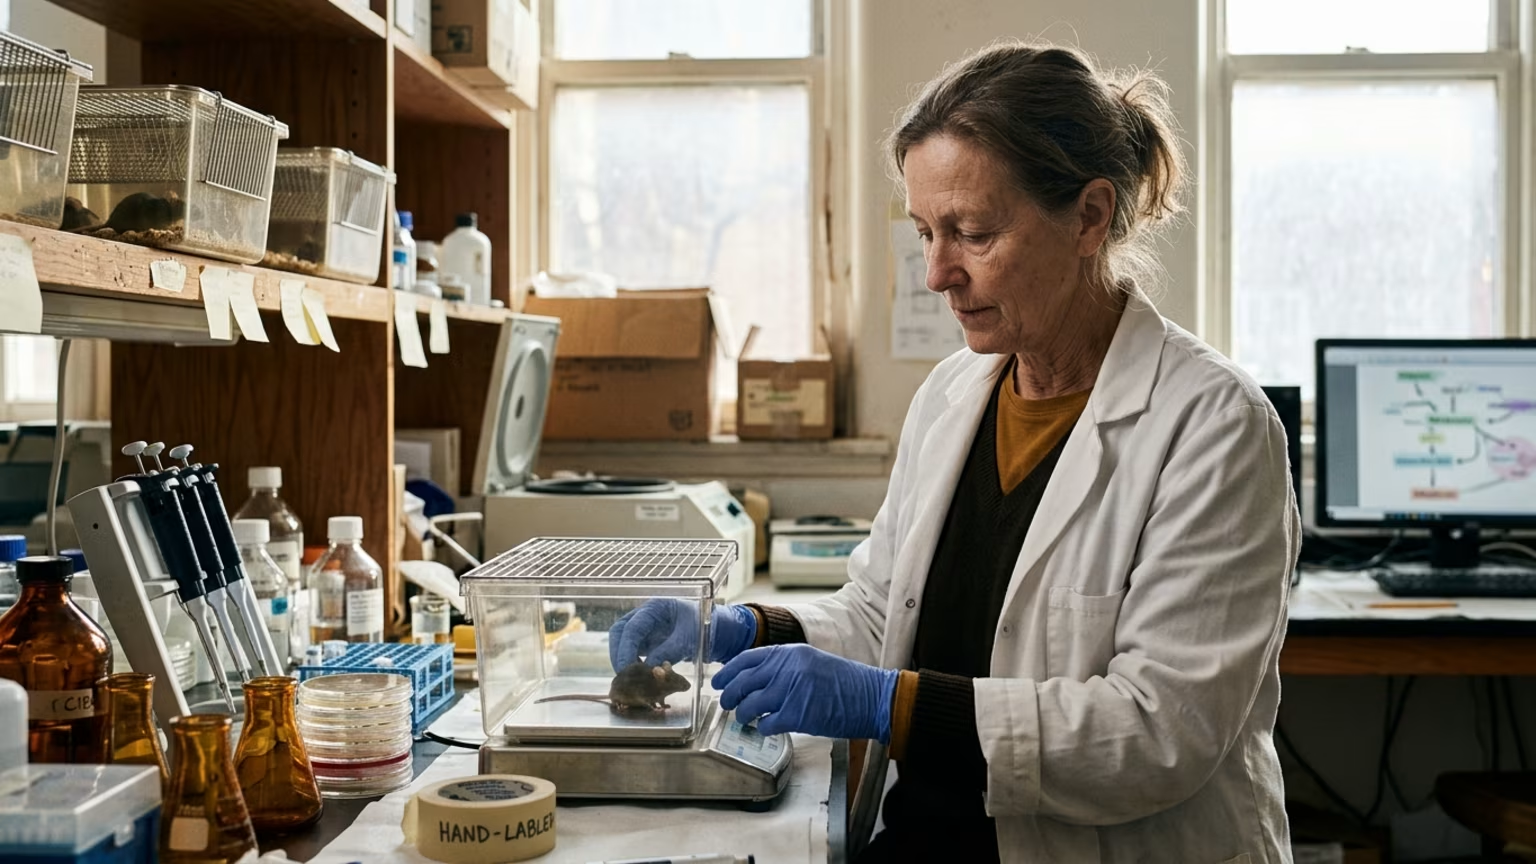
On July 29, 2026, the study “Mitochondrial metabolism and epigenetic crosstalk drive SASP” was reported as published in Nature. Researchers at Sanford Burnham Prebys worked with Mayo Clinic and collaborating institutions on the research.
Sanford Burnham Prebys reports that blocking a gene connected to mitochondrial metabolism reduced inflammation and promoted healthier aging in mice. The study links mitochondrial citrate metabolism to the accessibility of inflammatory genes in senescent cells. Its central finding is that metabolism helps regulate the senescence-associated secretory phenotype, or SASP, alongside immune signals associated with leaked mitochondrial genetic material. For longevity research, the result identifies a potential point of intervention between a cell’s metabolic activity and the sustained inflammatory molecules it releases.
Study setting: preclinical research in animal models, with the reported healthy-aging benefits observed in mice.
Senescent cells stop dividing but can remain active. According to the institutional account, the research examines the inflammatory molecules these cells release through the SASP. The question is not simply whether senescent cells are present. It is how mitochondrial activity helps regulate their inflammatory output.
A study-focused account of the mechanism describes a connection between mitochondrial citrate metabolism and changes in chromatin that affect inflammatory gene accessibility. Separate coverage describes mitochondrial DNA and RNA leakage as another contributor to immune signaling. These are linked parts of the proposed explanation rather than competing accounts of the same event.
This is a useful context for considering mitochondrial functions beyond energy production. Here, the research connects metabolism with the regulation of an inflammatory gene program. The biological question is therefore about the relationship between metabolic activity and inflammatory behavior, not simply whether mitochondria are producing enough energy.
In the reported model, increased export of citrate from mitochondria supports acetyl-CoA production. Acetyl-CoA availability is linked to changes involving histones and chromatin. Those changes can make SASP-related inflammatory genes more accessible.
The sequence matters because the study places a metabolic process upstream of inflammatory gene activity. Citrate export is not described merely as another measurement that changes in senescent cells. The study-focused report presents it as part of the pathway supporting the inflammatory program. That relationship is what makes the transport step an experimental target.
The distinction between gene accessibility and an inflammatory signal is central to interpreting the result. Making an inflammatory gene accessible is a different contribution from supplying the signal that activates an immune response. In this model, mitochondrial metabolism helps establish conditions in which the inflammatory program can operate.
This also explains the study title’s reference to metabolic and epigenetic crosstalk. The proposed connection runs from citrate metabolism through acetyl-CoA availability to chromatin changes and SASP-related gene expression. The result brings these processes into the same explanation of senescent-cell inflammation rather than treating metabolism and gene regulation as separate subjects.
The other part of the proposed mechanism involves mitochondrial nucleic acids. According to the available coverage, leaked mitochondrial DNA and RNA can activate immune signaling. Their role differs from the metabolic contribution to gene accessibility.
A useful way to read the model is to keep those contributions distinct. The citrate and acetyl-CoA pathway helps make inflammatory genes accessible. Mitochondrial nucleic-acid leakage supplies an inflammatory signal. The study brings these processes together to explain how senescent cells sustain inflammatory activity.
Coverage of the intervention reports that blocking the metabolic pathway reduced its contribution to inflammatory gene activity without removing the immune signals associated with mitochondrial DNA and RNA leakage. That makes the result more specific than a general claim that the experiment switched off inflammation. The reported intervention acted on one contribution to the inflammatory program while another remained present.
For readers comparing explanations of the study, this distinction helps reconcile apparently different descriptions. An account emphasizing gene accessibility and an account emphasizing immune signaling may be discussing different parts of the same proposed mechanism. The important question is which part the intervention actually changed.
The study-focused account identifies SLC25A1 as the mitochondrial citrate carrier involved in the pathway. Blocking this transport step was reported to reduce inflammatory features. The target therefore sits within the proposed connection between citrate export and acetyl-CoA availability.
The logic of the experiment is narrower than trying to suppress every source of inflammatory signaling. It tests whether interfering with a metabolic contribution changes the inflammatory program. Read this way, the study is an investigation of how the SASP is regulated as well as a report of an intervention’s effects.
The mouse observations add an organism-level result to that mechanistic account. Sanford Burnham Prebys describes reduced inflammation and healthier aging after blocking a related gene. The pathway finding and the animal finding belong together, but they answer different questions: one concerns how inflammatory gene activity is supported and the other concerns the consequences observed in mice.
Keeping those questions separate is useful when weighing the study. A coherent mechanism explains why an intervention was tested. The animal outcomes establish what was reported after that intervention. Neither should substitute for a precise account of the other.
The institutional summary and accompanying study coverage leave important questions unresolved. These are the boundaries to keep attached to the reported result:
When assessing future coverage, keep the intervention, the measured outcome and the organism together in the same claim. This makes it easier to recognize when a mechanistic finding has been stretched into a broader longevity conclusion. The useful reading task is to identify what changed in the experiment before assigning significance to the healthy-aging language.
The next useful phase is independent replication with defined healthspan outcomes and safety data, followed by human research designed to test whether targeting the same pathway produces a measurable clinical benefit.
Judging a citrate-blocking experiment means checking whether its inflammatory findings support its healthy-aging claims, and AgeAmaze helps readers make sense of that evidence. Its coverage addresses confusion between longevity studies by explaining mechanisms alongside experimental limits, giving skeptical adults a clearer basis for interpreting emerging interventions.
Stay current with research on aging biology, biomarkers, nutrition, therapeutics, peptides and longevity technology. AgeAmaze reports what the evidence shows, where uncertainty remains and which claims still need stronger data.




Follow AgeAmaze for careful reporting on what longevity science can show today and what still needs stronger evidence.
read the Blog